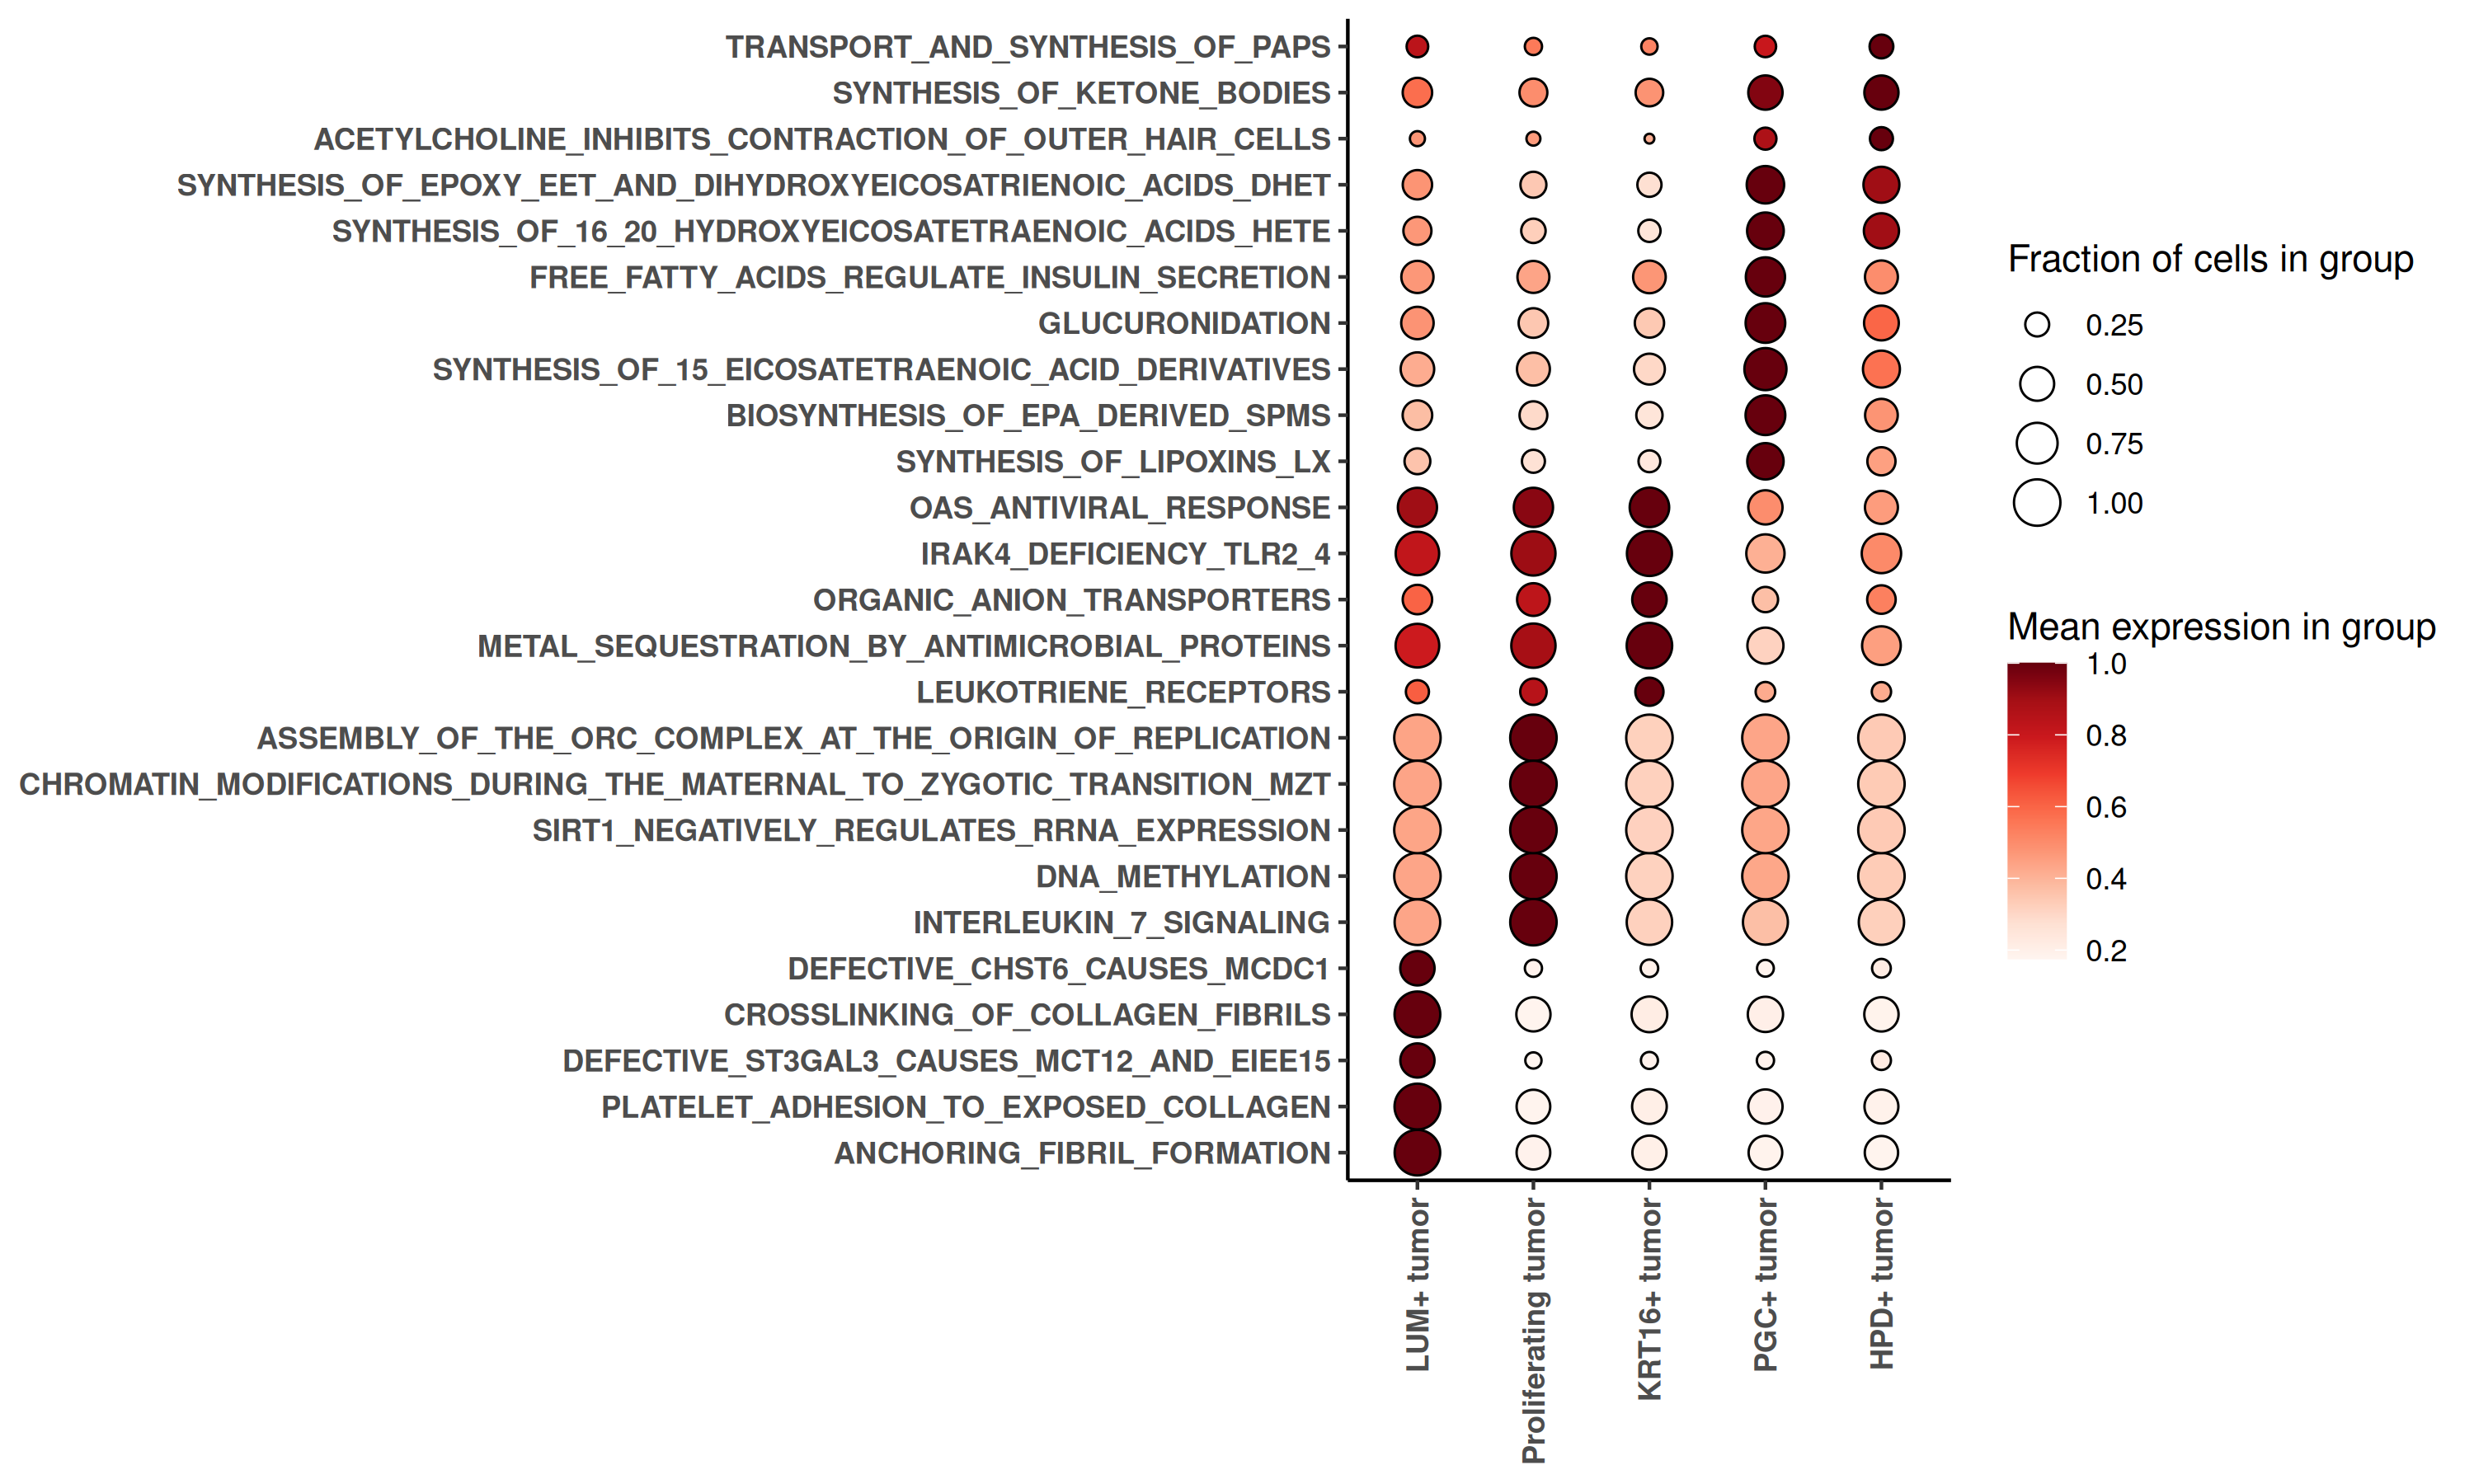
In this section we use AUCell to calculate pathway activity across thousands of pathways. We use pathways found in the Reactome database for this analysis.
AUCell operates independently on cells and on pathways, so we use chunking of our dataset to run all these calculations.
Show the code
# Read in gene sets and chunk for processing# original location: gene.sets <- readRDS("~/data/dan/imputation_smoothing/breast_reanalysis/gene.sets.list.rds")[["human"]]gene.sets <-readRDS(paste0(pathway_dir, "/Data/gene.sets.list.rds"))[["human"]]chunks <-data.table(strt=unique(c(seq(1, length(gene.sets), 100), length(gene.sets))))chunks[,stp:=data.table::shift(strt, 1, type="lead")]chunks <- chunks[-c(.N)]chunks[1:(.N-1),stp:=stp-1]# if there's no seurat object, read it inif (!("seu.obj"%in%ls())){ seu.obj <-readRDS(results_list$seurat_file_postCellTyping)}# Extract normalized data and meta datatc <- Matrix::colSums(seu.obj[["RNA"]]@counts)scale.factor <-mean(tc)normed <- seu.obj[["RNA"]]@counts %*% Matrix::Diagonal(x = scale.factor / tc, names =TRUE)met <-data.table(seu.obj@meta.data)# Calculate median diversity percentilemedian_diversity_pctile <-median(met$nFeature_RNA) /nrow(normed)################################################################################ Run AUCell by chunks of cells##############################################################################cells <- met[["cell_ID"]]cellchunks <-data.table(cells)[,chunk:=floor((1:.N)/20000) +1][]for(ii in1:cellchunks[,max(chunk)]){message(ii) cellsscore <- cellchunks[chunk==ii][["cells"]] chunk_counts <- normed[,cellsscore]set.seed(250404) cells_rankings <- AUCell::AUCell_buildRankings(exprMat=as(chunk_counts, "dgCMatrix"), ,plotStats =FALSE ,splitByBlocks =TRUE ,BPPARAM = BiocParallel::MulticoreParam(workers =3) )saveRDS(cells_rankings, paste0(pathway_dir, "/Data/cells_ranking_cellchunk_", ii, ".rds")) pb <-txtProgressBar(min =0, max =nrow(chunks), style=3) for(chunky in1:nrow(chunks)){message(chunky) strt <- chunks[chunky,strt] stp <- chunks[chunky,stp]#run aucell for all gene sets reschunk <- AUCell::AUCell_calcAUC(geneSets=gene.sets[strt:stp], cells_rankings, aucMaxRank=ceiling(nrow(cells_rankings)*median_diversity_pctile), verbose =TRUE,nCores =4, ) reschunk <-as(reschunk@assays@data$AUC, "sparseMatrix")setTxtProgressBar(pb, chunky)saveRDS(reschunk, file=paste0(pathway_dir, "/Data/cellsAUC_" ,".cellschunk__",ii ,".chunk__",chunky ,".rds") )rm(reschunk); gc() } gc()}# Join all cells and pathways to one matrixcells_auc_full <-NULLfor(ii in1:cellchunks[,max(chunk)]){ cells_auc_mtx <-readRDS(file=paste0(pathway_dir, "/Data/cellsAUC_" ,".cellschunk__",ii ,".chunk__","1" ,".rds"))for(chunky in2:nrow(chunks)){ temp_mtx <-readRDS(file=paste0(pathway_dir, "/Data/cellsAUC_" ,".cellschunk__",ii ,".chunk__",chunky ,".rds")) cells_auc_mtx <-rbind(cells_auc_mtx, temp_mtx) }if(is.null(cells_auc_full)){ cells_auc_full <- cells_auc_mtx } else{ cells_auc_full <-cbind(cells_auc_full, cells_auc_mtx) }}saveRDS(cells_auc_full, file =paste0(pathway_dir, "/Data/cellsAUC_" ,"full" ,".rds"))
Make a heatmap of the top pathways by tumor subtype.
Show the code
# if there's no seurat object, read it inif (!("seu.obj"%in%ls())){ seu.obj <-readRDS(results_list$seurat_file_postCellTyping)}met =as.data.table(seu.obj@meta.data)# Read in cells x pathways score matrix, pathway data (pwd)pwd <-readRDS(file =paste0(pathway_dir, "/Data/cellsAUC_" ,"full" ,".rds"))## Heatmap of top differentiating pathways by tumor typetum_cells <- seu.obj$cell_id[grep("tumor", seu.obj$minor_cell_type)]pwfctbl_tum <- smiSmooth::clusterwise_foldchange_metrics(normed = pwd[,tum_cells] ,metadata = met[cell_ID %in% tum_cells,.(minor_cell_type, cell_ID)] ,cluster_column ="minor_cell_type")# Save itsaveRDS(pwfctbl_tum, file =paste0(pathway_dir, "/Data/pwfctbl_tum.rds"))# Change pathway namespwfctbl_tum$gene <-gsub("REACTOME_", "", pwfctbl_tum$gene)# Make a heatmap showing top differentiating pathway by cell typepng(filename =paste0(pathway_dir, "/Plots/heatmap_pathways_by_cell_type_tumors.png"),height =6, width =10, units ="in", res =300)smiSmooth::marker_heatmap(pwfctbl_tum, topn=5) +coord_flip()dev.off()